O problema com as "teorias de multiversos": elas simplesmente não são científicas!!!

terça-feira, julho 10, 2018

The problem with “multiverse theories”: they’re just not science

This admittedly catchy idea is undermining the integrity of physics

by Jim Baggott / June 25, 2018 /


Over the last few decades “multiverse theories” have become increasingly fashionable within a relatively small—but publicly vocal—group of theoretical physicists. This group specialises in foundational problems in cosmology, particle physics, and quantum mechanics. These theories are advertised as science’s answer to much that we can’t otherwise explain about the universe we inhabit, the elementary particles we have discovered in it, and the reasons for our own existence.


It’s “theories” plural because the multiverse is used in various ways to fill gaps in our current understanding. Cosmological multiverse theories “explain” why the initial conditions that prevailed at the Big Bang origin of our universe, and the physical constants and laws which shaped its subsequent evolution, appear so exquisitely fine-tuned to allow for the possibility of life. The idea is that there’s nothing particularly special about our “Goldilocks” universe: it is simply one of a (possibly infinite) number of universes, all with different initial conditions, constants, and laws. Most will be inhospitable, but it should come as no surprise to find ourselves in a universe which isn’t.

...


Read more here: Prospect Magazine

Darwin, mais design inteligente, mano: sistema de encriptação descoberto em genes!!!

segunda-feira, julho 09, 2018

Transient N-6-Methyladenosine Transcriptome Sequencing Reveals a Regulatory Role of m6A in Splicing Efficiency

Annita Louloupi5, Evgenia Ntini5, Thomas Conrad, Ulf Andersson Vang Ørom6

5These authors contributed equally

6Lead Contact


Article Info

Publication History

Published: June 19, 2018 Accepted: May 23, 2018

Received in revised form: April 30, 2018 Received: January 10, 2018

User License

Creative Commons Attribution – NonCommercial – NoDerivs (CC BY-NC-ND 4.0)

Source/Fonte: TeskaLabs

Highlights

•A time-resolved high-resolution picture of m6A on nascent RNA transcripts

•m6A is deposited at nascent RNA and in introns

•m6A deposition at splice-junctions increases splicing kinetics

•High m6A levels in introns is associated with slow and alternative splicing

Summary

Splicing efficiency varies among transcripts, and tight control of splicing kinetics is crucial for coordinated gene expression. N-6-methyladenosine (m6A) is the most abundant RNA modification and is involved in regulation of RNA biogenesis and function. The impact of m6A on regulation of RNA splicing kinetics is unknown. Here, we provide a time-resolved high-resolution assessment of m6A on nascent RNA transcripts and unveil its importance for the control of RNA splicing kinetics. We find that early co-transcriptional m6A deposition near splice junctions promotes fast splicing, while m6A modifications in introns are associated with long, slowly processed introns and alternative splicing events. In conclusion, we show that early m6A deposition specifies the fate of transcripts regarding splicing kinetics and alternative splicing.

Ainda por explicar: quase um em cada cinco genes humanos ainda tem status de codificação não resolvido

Loose ends: almost one in five human genes still have unresolved coding status 

Federico Abascal David Juan Irwin Jungreis Laura Martinez Maria Rigau Jose Manuel Rodriguez Jesus Vazquez Michael L Tress

Nucleic Acids Research, gky587, https://doi.org/10.1093/nar/gky587

Published: 30 June 2018 

Article history Received: 15 May 2018 Revision Received: 12 June 2018

Accepted: 18 June 2018


Abstract

Seventeen years after the sequencing of the human genome, the human proteome is still under revision. One in eight of the 22 210 coding genes listed by the Ensembl/GENCODE, RefSeq and UniProtKB reference databases are annotated differently across the three sets. We have carried out an in-depth investigation on the 2764 genes classified as coding by one or more sets of manual curators and not coding by others. Data from large-scale genetic variation analyses suggests that most are not under protein-like purifying selection and so are unlikely to code for functional proteins. A further 1470 genes annotated as coding in all three reference sets have characteristics that are typical of non-coding genes or pseudogenes. These potential non-coding genes also appear to be undergoing neutral evolution and have considerably less supporting transcript and protein evidence than other coding genes. We believe that the three reference databases currently overestimate the number of human coding genes by at least 2000, complicating and adding noise to large-scale biomedical experiments. Determining which potential non-coding genes do not code for proteins is a difficult but vitally important task since the human reference proteome is a fundamental pillar of most basic research and supports almost all large-scale biomedical projects.

Issue Section: Data Resources and Analyses

FREE PDF GRATIS: Nucleic Acids Research

Análise comparativa de alta resolução de genomas de grandes primatas

High-resolution comparative analysis of great ape genomes

Zev N. Kronenberg1, Ian T. Fiddes2,*, David Gordon1,3,*, Shwetha Murali1,3,*, Stuart Cantsilieris1,*, Olivia S. Meyerson4,*, Jason G. Underwood1,5,*, Bradley J. Nelson1,*, Mark J. P. Chaisson1,6, Max L. Dougherty1, Katherine M. Munson1, Alex R. Hastie7, Mark Diekhans2, Fereydoun Hormozdiari8, Nicola Lorusso9, Kendra Hoekzema1, Ruolan Qiu1, Karen Clark10, Archana Raja1,3, AnneMarie E. Welch1, Melanie Sorensen1, Carl Baker1, Robert S. Fulton11, Joel Armstrong2, Tina A. Graves-Lindsay11, Ahmet M. Denli12, Emma R. Hoppe1, PingHsun Hsieh1, Christopher M. Hill1, Andy Wing Chun Pang7, Joyce Lee7, Ernest T. Lam7, Susan K. Dutcher11, Fred H. Gage12, Wesley C. Warren11, Jay Shendure1,3, David Haussler2,13, Valerie A. Schneider10, Han Cao7, Mario Ventura9, Richard K. Wilson11, Benedict Paten2, Alex Pollen4,14, Evan E. Eichler1,3,†

See all authors and affiliations

Science 08 Jun 2018: Vol. 360, Issue 6393, eaar6343


A spotlight on great ape genomes

Most nonhuman primate genomes generated to date have been “humanized” owing to their many gaps and the reliance on guidance by the reference human genome. To remove this humanizing effect, Kronenberg et al. generated and assembled long-read genomes of a chimpanzee, an orangutan, and two humans and compared them with a previously generated gorilla genome. This analysis recognized genomic structural variation specific to humans and particular ape lineages. Comparisons between human and chimpanzee cerebral organoids showed down-regulation of the expression of specific genes in humans, relative to chimpanzees, related to noncoding variation identified in this analysis.

Science, this issue p. eaar6343

Structured Abstract

INTRODUCTION

Understanding the genetic differences that make us human is a long-standing endeavor that requires the comprehensive discovery and comparison of all forms of genetic variation within great ape lineages.


RATIONALE

The varied quality and completeness of ape genomes have limited comparative genetic analyses. To eliminate this contiguity and quality disparity, we generated human and nonhuman ape genome assemblies without the guidance of the human reference genome. These new genome assemblies enable both coarse and fine-scale comparative genomic studies.


RESULTS

We sequenced and assembled two human, one chimpanzee, and one orangutan genome using high-coverage (>65x) single-molecule, real-time (SMRT) long-read sequencing technology. We also sequenced more than 500,000 full-length complementary DNA samples from induced pluripotent stem cells to construct de novo gene models, increasing our knowledge of transcript diversity in each ape lineage. The new nonhuman ape genome assemblies improve gene annotation and genomic contiguity (by 30- to 500-fold), resulting in the identification of larger synteny blocks (by 22- to 74-fold) when compared to earlier assemblies. Including the latest gorilla genome, we now estimate that 83% of the ape genomes can be compared in a multiple sequence alignment.

We observe a modest increase in single-nucleotide variant divergence compared to previous genome analyses and estimate that 36% of human autosomal DNA is subject to incomplete lineage sorting. We fully resolve most common repeat differences, including full-length retrotransposons such as the African ape-specific endogenous retroviral element PtERV1. We show that the spread of this element independently in the gorilla and chimpanzee lineage likely resulted from a founder element that failed to segregate to the human lineage because of incomplete lineage sorting.

The improved sequence contiguity allowed a more systematic discovery of structural variation (>50 base pairs in length) (see the figure). We detected 614,186 ape deletions, insertions, and inversions, assigning each to specific ape lineages. Unbiased genome scaffolding (optical maps, bacterial artificial chromosome sequencing, and fluorescence in situ hybridization) led to the discovery of large, unknown complex inversions in gene-rich regions. Of the 17,789 fixed human-specific insertions and deletions, we focus on those of potential functional effect. We identify 90 that are predicted to disrupt genes and an additional 643 that likely affect regulatory regions, more than doubling the number of human-specific deletions that remove regulatory sequence in the human lineage. We investigate the association of structural variation with changes in human-chimpanzee brain gene expression using cerebral organoids as a proxy for expression differences. Genes associated with fixed structural variants (SVs) show a pattern of down-regulation in human radial glial neural progenitors, whereas human-specific duplications are associated with up-regulated genes in human radial glial and excitatory neurons (see the figure).

CONCLUSION

The improved ape genome assemblies provide the most comprehensive view to date of intermediate-size structural variation and highlight several dozen genes associated with structural variation and brain-expression differences between humans and chimpanzees. These new references will provide a stepping stone for the completion of great ape genomes at a quality commensurate with the human reference genome and, ultimately, an understanding of the genetic differences that make us human.

Abstract

Genetic studies of human evolution require high-quality contiguous ape genome assemblies that are not guided by the human reference. We coupled long-read sequence assembly and full-length complementary DNA sequencing with a multiplatform scaffolding approach to produce ab initio chimpanzee and orangutan genome assemblies. By comparing these with two long-read de novo human genome assemblies and a gorilla genome assembly, we characterized lineage-specific and shared great ape genetic variation ranging from single– to mega–base pair–sized variants. We identified ~17,000 fixed human-specific structural variants identifying genic and putative regulatory changes that have emerged in humans since divergence from nonhuman apes. Interestingly, these variants are enriched near genes that are down-regulated in human compared to chimpanzee cerebral organoids, particularly in cells analogous to radial glial neural progenitors.


SUBSCRIPTION OR PAYMENT NEEDED/REQUER ASSINATURA OU PAGAMENTO:

Science

+++++

Professores, pesquisadores e alunos de universidades públicas e privadas com acesso ao Portal Periódicos CAPES/MEC podem ler gratuitamente este artigo da Science e mais 33.000 publicações científicas.

Distribuição desigual da variância mutacional em todo o transcriptoma da Drosophila serrata revelada pela análise de alta dimensionalidade da expressão gênica

Uneven Distribution of Mutational Variance Across the Transcriptome of Drosophila serrata Revealed by High-Dimensional Analysis of Gene Expression

Emma Hine, Daniel E. Runcie, Katrina McGuigan and Mark W. Blows

GENETICS Early online June 8, 2018;


Abstract

There are essentially an infinite number of traits that could be measured on any organism, and almost all individual traits display genetic variation, yet substantial genetic variance in a large number of independent traits is not plausible under basic models of selection and mutation. One mechanism that may be invoked to explain the observed levels of genetic variance in individual traits is that pleiotropy results in fewer dimensions of phenotypic space with substantial genetic variance. Multivariate genetic analyses of small sets of functionally-related traits have shown that standing genetic variance is often concentrated in relatively few dimensions. It is unknown if a similar concentration of genetic variance occurs at a phenome-wide scale when many traits of disparate function are considered, or if the genetic variance generated by new mutations is also unevenly distributed across phenotypic space. Here, we used a Bayesian sparse factor model to characterize the distribution of mutational variance of 3385 gene expression traits of Drosophila serrata after 27 generations of mutation accumulation, and found that 46% of the estimated mutational variance was concentrated in just 21 dimensions with significant mutational heritability. We show that the extent of concentration of mutational variance into such a small subspace has the potential to substantially bias the response to selection of these traits.

GENETIC CONSTRAINTS GENE EXPRESSION MUTATION PLEIOTROPY

Received May 10, 2018. Revision received May 10, 2018.

Accepted May 31, 2018.

Copyright © 2018, Genetics

FREE PDF GRATIS: Genetics Sup. Info.

Entendendo o progresso da ciência

McCoy, C.D. (2018) Understanding the Progress of Science. [Preprint]


Abstract

This paper develops a problem-solving account of scientific progress that takes understanding as the principal epistemic aim of science. It examines a recent paper of Bird's on scientific progress, argues for the primacy of understanding over knowledge in this context, illustrates the account using a Kuhnian picture of science, and defends it against knowledge reductionism.

FREE PDF GRATIS: PhilSci

Rastros ediacaranos tardios produzidos por animais bilatérios com apêndices pareados


Late Ediacaran trackways produced by bilaterian animals with paired appendages

Zhe Chen1,2,*, Xiang Chen1,3, Chuanming Zhou2,4, Xunlai Yuan1,2,3 and Shuhai Xiao5,*

1State Key Laboratory of Palaeobiology and Stratigraphy, Nanjing Institute of Geology and Palaeontology, Chinese Academy of Sciences, Nanjing 210008, China.

2Center for Excellence in Life and Paleoenvironment, Chinese Academy of Sciences, Nanjing 210008, China.

3University of Chinese Academy of Sciences, Beijing 100049, China.

4CAS Key Laboratory of Economic Stratigraphy and Palaeogeography, Nanjing Institute of Geology and Palaeontology, Chinese Academy of Sciences, Nanjing 210008, China.

5Department of Geosciences, Virginia Polytechnic Institute and State University, Blacksburg, VA 24061, USA.

↵*Corresponding author. Email: xiao@vt.edu (S.X.); zhechen@nigpas.ac.cn (Z.C.)

Science Advances 06 Jun 2018:

Vol. 4, no. 6, eaao6691


Trackways and burrows excavated in situ from the Shibantan Member.

Abstract

Ediacaran trace fossils provide key paleontological evidence for the evolution of early animals and their behaviors. Thus far, however, this fossil record has been limited to simple surface trails and relatively shallow burrows. We report possible trackways, preserved in association with burrows, from the terminal Ediacaran Shibantan Member (ca. 551 to ca. 541 million years ago) in the Yangtze Gorges area of South China. These trace fossils represent the earliest known trackways. They consist of two rows of imprints arranged in poorly organized series or repeated groups. These trackways may have been produced by bilaterian animals with paired appendages, although the phylum-level phylogenetic affinity of the trace makers remains unknown. It is possible that the trackways and associated burrows were produced by the same trace maker, indicating a complex behavior involving both walking and burrowing. Together, these trackways and burrows mark the arrival of a new era characterized by an increasing geobiological footprint of bilaterian animals.

FREE PDF GRATIS: Science Advances Sup. Info.

Nova dinâmica inesperada para grandes moléculas de DNA em suspensão líquida

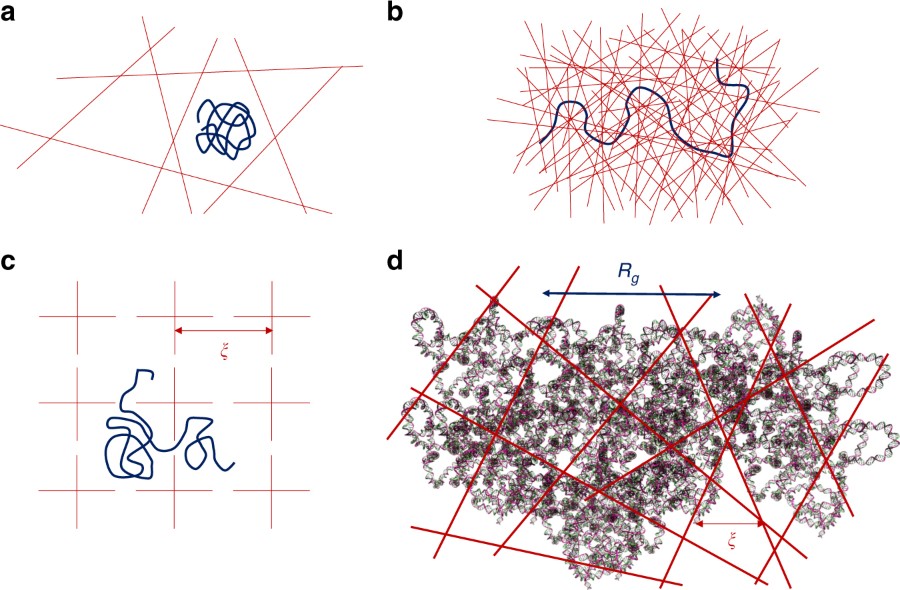
Topologically frustrated dynamics of crowded charged macromolecules in charged hydrogels

Di Jia & Murugappan Muthukumar 

Nature Communications volume 9, Article number: 2248 (2018) 

Fig. 1

Abstract

Movement of charged macromolecules in crowded aqueous environments is a ubiquitous phenomenon vital to the various living processes and formulations of materials for health care. While study of diffusion of tracer amounts of probe macromolecules trapped inside concentrated solutions, gels, or random media has led to an enhanced understanding of this complex process, the collective dynamics of charged macromolecules embedded inside congested charge-bearing matrices still remains to be fully explored. Here we report a frustrated dynamics of DNA and synthetic polyelectrolytes inside a charged host hydrogel where the guest molecules do not diffuse. Instead, they exhibit a family of relaxation processes arising from a combination of conformational entropy and local chain dynamics, which are frustrated by the confinement from the gel. We also have developed a model explaining this new universality class of non-diffusive topologically frustrated dynamics of charged macromolecules.

Acknowledgements

We thank the National Science Foundation (DMR-1504265), AFOSR Grant FA9550-17-1-0160, and the National Institutes of Health (Grant No. R01HG002776-11) for financial support.

Author information

Affiliations

Department of Polymer Science and Engineering, University of Massachusetts, Amherst, MA, 01003, USA

Di Jia & Murugappan Muthukumar

Contributions

M.M. conceived the project. D.J. designed the experiments, synthesized the gel, and performed all experiments. M.M. developed the theory. Both authors discussed and contributed to the interpretation of the data. The whole project was supervised by M.M.

Competing interests

The authors declare no competing interests.

Corresponding author

Correspondence to Murugappan Muthukumar.

O espectro de muitas espécies

domingo, julho 08, 2018

2018 Jul Syst Biol - Leaché et al - The Spectre of Too Many Species

Adam D Leaché Tianqi Zhu Bruce Rannala Ziheng Yang

Systematic Biology, syy051, https://doi.org/10.1093/sysbio/syy051

Published: 05 July 2018 Article history

Received: 07 November 2017 Revision Received: 22 June 2018

Accepted: 29 June 2018


Abstract

Recent simulation studies examining the performance of Bayesian species delimitation as implemented in the BPP program have suggested that BPP may detect population splits but not species divergences and that it tends to over-split when data of many loci are analyzed. Here we confirm these results and provide the mathematical justifications. We point out that the distinction between population and species splits made in the protracted speciation model has no influence on the generation of gene trees and sequence data, which explains why no method can use such data to distinguish between population splits and speciation. We suggest that the protracted speciation model is unrealistic as its mechanism for assigning species status assumes instantaneous speciation, contradicting prevailing taxonomic practice. We confirm the suggestion, based on simulation, that in the case of speciation with gene flow, Bayesian model selection as implemented in BPP tends to detect population splits when the amount of data (the number of loci) increases. We discuss the use of a recently proposed empirical genealogical divergence index (gdi) for species delimitation and illustrate that parameter estimates produced by a full likelihood analysis as implemented in BPP provide much more reliable inference under the gdi than the approximate method PHRAPL. We distinguish between Bayesian model selection and parameter estimation, and suggest that the model selection approach is useful for identifying sympatric cryptic species while the parameter estimation approach may be used to implement empirical criteria for determining species status among allopatric populations.

Species delimitation, BPP, multispecies coalescent, taxonomy

Issue Section: POINT OF VIEW

© The Author(s) 2018. Published by Oxford University Press, on behalf of the Society of Systematic Biologists.

This is an Open Access article distributed under the terms of the Creative Commons Attribution Non-Commercial License (http://creativecommons.org/licenses/by-nc/4.0/), which permits non-commercial re-use, distribution, and reproduction in any medium, provided the original work is properly cited. For commercial re-use, please contact journals.permissions@oup.com

FREE PDF GRATIS: Systematic Biology

As discrepâncias de dados podem afetar nosso entendimento do universo

Cosmological discordances. II. Hubble constant, Planck and large-scale-structure data sets

Weikang Lin and Mustapha Ishak

Phys. Rev. D 96, 083532 – Published 30 October 2017


A visual history of the expanding Universe includes the hot, dense state known as the Big Bang and the growth and formation of structure subsequently. But quantitatively knowing what the expansion rate is (and was) in the present (and past) is vital to understanding our cosmic history and future. Source/Fonte

ABSTRACT

We examine systematically the (in)consistency between cosmological constraints as obtained from various current data sets of the expansion history, large-scale-structure (LSS), and cosmic microwave background (CMB) temperature and polarization from Planck. We run (dis)concordance tests within each set and across the three sets using a recently introduced index of inconsistency (IOI) capable of dissecting inconsistencies between two or more data sets. First, we compare the constraints on 
H0
 from five different methods and find that the IOI drops from 2.85 to 0.88 (on Jeffreys’s scales) when the local 
H0
 measurements are removed. This seems to indicate that the local measurement is an outlier compared to the others, thus favoring a systematics-based explanation. We find a moderate inconsistency (
IOI=2.61
) between Planck temperature and polarization data sets. We find that current LSS data sets including the WiggleZ power spectrum, SDSS redshift space distortion, CFHTLenS weak lensing, CMB lensing, and cluster count from SZ effect, are consistent one with another and also when all combined. However, we find a persistent moderate inconsistency between Planck and individual or combined LSS probes. For Planck 
TT+lowTEB
 versus individual LSS probes, the IOI spans the range 2.92–3.72 and increases to 3.44–4.20 when the polarization data is added in. The joint LSS versus the combined Planck temperature and polarization has an IOI of 2.83 in the most conservative case. But if Planck low-
ℓ
 temperature and polarization is also added to the joint LSS to constrain 
τ
 and break degeneracies, the inconsistency between Planck and joint LSS data increases to the high end of the moderate range with 
IOI=4.81
. Whether due to systematic effects in the data or to the underlying model, these inconsistencies need to be resolved. Finally, we perform forecast calculations using the Large Sky Synoptic Survey (LSST) and find that the discordance between Planck and future LSS data, if it persists as present, can rise up to a high IOI of 17, thus falling in the strong range of inconsistency.


Received 1 September 2017
© 2017 American Physical Society
Physics Subject Headings (PhySH)
Research Areas Cosmological parameters Cosmology Gravitation, Cosmology & Astrophysics
Subscription or payment needed/Requer assinatura ou pagamento:

Physical Review D 

Francisco Ayala, famoso geneticista evolucionista, perde cargo: assédio sexual contra várias mulheres

terça-feira, julho 03, 2018

Acclaimed UC Irvine geneticist who gave millions to the campus resigns due to sexual harassment

By Teresa Watanabe


Acclaimed UC Irvine geneticist Francisco J. Ayala has resigned after a university investigation found he sexually harassed four faculty members and graduate students, the university has announced.

Ayala, a former Dominican priest who won the prestigious Templeton Prize for championing both science and faith, resigned effective July 1 and will abstain from future university activities, UCI officials said.

“I deeply regret that what I have always thought of as the good manners of a European gentleman — to greet women colleagues warmly, with a kiss to both cheeks, to compliment them on their beauty — made colleagues I respect uncomfortable,” Ayala, who grew up in Madrid, said Friday in a statement. “It was never my intent to do so.”

He said he had “too much respect” for the women, his family and UC Irvine to continue defending himself with hearings, appeals or lawsuits and would continue his research “with renewed vigor” elsewhere.

In 2011, Ayala donated $10 million to the School of Biological Sciences, which then bore his name. It was the largest gift from a faculty member at the time.

The university said Ayala’s name has been removed from that school, and also is being removed from its central science library, graduate fellowships, scholar programs and endowed chairs. The biology school will now be known as the UCI School of Biological Sciences.

“I thank and commend our colleagues who reported this misconduct,” Chancellor Howard Gillman said in a statement Thursday. “Coming forward with this information was extremely courageous. Professor Ayala’s behavior defied our core beliefs and was inconsistent with our policies, guidelines and required training.”

Micha Liberty, an attorney who represents three of the women, said UCI ignored years of complaints from professors and graduate students that Ayala touched them and made sexual and sexist comments. She said one of the professors she’s representing reported Ayala’s conduct three years ago, but university officials failed to investigate or sanction him.

“They just told him, ‘Stay away from her,’ ” Liberty said. “Dr. Ayala has had a long and successful career and was clearly an asset to the UCI campus … and that in turn motivated UCI to look the other way when it came to complaints of sexual harassment.”

The university started the investigation last November. The women, who asked to be identified, are Kathleen Treseder, professor and chair of ecology and evolutionary biology; Jessica Pratt, an assistant teaching professor; Benedicte Shipley, an assistant dean; and Michelle Herrera, a graduate student.

After interviews with the women and more than 60 witnesses, the university substantiated the complaints last month.

UCI advisors are reaching out to the school’s faculty, students, staff and administration to provide counseling services and harassment-related education, officials said.

Liberty said her clients are dissatisfied that UCI has not acknowledged its failure to act on previous complaints and protect women from Ayala. She said there are many more victims but most are scared to come forward because his stature in the field gives him the power to make or break careers.

“In the face of overwhelming evidence, UCI came up with an anemic response to a systemic failure,” she said.
...
Read more here: Los Angeles Times

Darwin, o biofísico Ken Dill 'falou e disse': as máquinas de proteínas são MÁQUINAS, não são METÁFORAS!!!

quinta-feira, junho 28, 2018

Alô ETs, disquem 0800-NASA

terça-feira, junho 26, 2018

Exoplanet Biosignatures: A Review of Remotely Detectable Signs of Life

Schwieterman Edward W. , Kiang Nancy Y. , Parenteau Mary N. , Harman Chester E. , DasSarma Shiladitya , Fisher Theresa M. , Arney Giada N. , Hartnett Hilairy E. , Reinhard Christopher T. , Olson Stephanie L. , Meadows Victoria S. , Cockell Charles S. , Walker Sara I. , Grenfell John Lee , … Show all Authors 

Published Online:1 Jun 2018 https://doi.org/10.1089/ast.2017.1729

Image result for ET
Source/Fonte: MovieWeb

Abstract

In the coming years and decades, advanced space- and ground-based observatories will allow an unprecedented opportunity to probe the atmospheres and surfaces of potentially habitable exoplanets for signatures of life. Life on Earth, through its gaseous products and reflectance and scattering properties, has left its fingerprint on the spectrum of our planet. Aided by the universality of the laws of physics and chemistry, we turn to Earth's biosphere, both in the present and through geologic time, for analog signatures that will aid in the search for life elsewhere. Considering the insights gained from modern and ancient Earth, and the broader array of hypothetical exoplanet possibilities, we have compiled a comprehensive overview of our current understanding of potential exoplanet biosignatures, including gaseous, surface, and temporal biosignatures. We additionally survey biogenic spectral features that are well known in the specialist literature but have not yet been robustly vetted in the context of exoplanet biosignatures. We briefly review advances in assessing biosignature plausibility, including novel methods for determining chemical disequilibrium from remotely obtainable data and assessment tools for determining the minimum biomass required to maintain short-lived biogenic gases as atmospheric signatures. We focus particularly on advances made since the seminal review by Des Marais et al. The purpose of this work is not to propose new biosignature strategies, a goal left to companion articles in this series, but to review the current literature, draw meaningful connections between seemingly disparate areas, and clear the way for a path forward. 

Key Words: Exoplanets—Biosignatures—Habitability markers—Photosynthesis—Planetary surfaces—Atmospheres—Spectroscopy—Cryptic biospheres—False positives. Astrobiology 18, 663–708.

FREE PDF GRATIS: Astrobiology

A vida, o universo, e tudo - 42 questões fundamentais

segunda-feira, junho 25, 2018

Life, the Universe, and everything—42 fundamental questions

Roland E Allen 1 and Suzy Lidström 2,3

Published 15 November 2016 • © 2016 The Royal Swedish Academy of Sciences

Physica Scripta, Volume 92, Number 1 


Author affiliations

1 Department of Physics and Astronomy, Texas A&M University College Station, Texas 77843, USA

2 Department of Physics and Astronomy, Uppsala University SE-75120 Uppsala, Sweden

3 Physica Scripta, Royal Swedish Academy of Sciences SE-104 05 Stockholm, Sweden

Dates

Received 5 August 2016 Accepted 24 October 2016 

Published 15 November 2016

Check for updates using Crossmark

Peer review information

Method: Single-blind

Revisions: 1

Screened for originality? Yes

Citation

Roland E Allen and Suzy Lidström 2017 Phys. Scr. 92 012501


Image result for questions marks

Abstract

In The Hitchhiker's Guide to the Galaxy, by Douglas Adams, the Answer to the Ultimate Question of Life, the Universe, and Everything is found to be 42—but the meaning of this is left open to interpretation. We take it to mean that there are 42 fundamental questions which must be answered on the road to full enlightenment, and we attempt a first draft (or personal selection) of these ultimate questions, on topics ranging from the cosmological constant and origin of the Universe to the origin of life and consciousness.

+++++

Original content from this work may be used under the terms of the Creative Commons Attribution 3.0 licence. Any further distribution of this work must maintain attribution to the author(s) and the title of the work, journal citation and DOI.

FREE PDF GRATIS: Physica Scripta

Dan Graur, o DNA "Lixo" é CRUCIAL no desenvolvimento de embriões!!!

sexta-feira, junho 22, 2018

A LINE1-Nucleolin Partnership Regulates Early Development and ESC Identity

Michelle Percharde, Chih-Jen Lin, Yafei Yin, Juan Guan, Gabriel A. Peixoto, Aydan Bulut-Karslioglu 6, Steffen Biechele, Bo Huang, Xiaohua Shen, Miguel Ramalho-Santos 7, 8

6 Present address: Max Planck Institute for Molecular Genetics, Berlin, Germany

7 Present address: Lunenfeld-Tanenbaum Research Institute and Department of Molecular Genetics, University of Toronto, Toronto, ON M5T 3H7, Canada

8 Lead Contact

Publication stage: In Press Corrected Proof

PlumX Metrics

Learn more about article metrics


Article Info

Publication History

Published: June 21, 2018 Accepted: May 17, 2018

Received in revised form: March 20, 2018 Received: August 12, 2017

In these two two-cell mouse embryos, the surface of the embryos is outlined in orange, the DNA in the nucleus is indicated in blue and the activity of the LINE-1 gene is indicated via bright red spots.
Ramalho-Santos lab/UCSF - Source/Fonte


Highlights

• LINE1 RNA is abundant and nuclear in mouse ESCs and pre-implantation embryos

• LINE1 knockdown inhibits ESC self-renewal and induces transition to a 2C state

• LINE1 RNA recruits Nucleolin/Kap1 to repress Dux and activate rRNA synthesis

• In embryos, LINE1 inhibition causes persistence of the 2C program and impairs ZGA

Summary

Transposable elements represent nearly half of mammalian genomes and are generally described as parasites, or “junk DNA.” The LINE1 retrotransposon is the most abundant class and is thought to be deleterious for cells, yet it is paradoxically highly expressed during early development. Here, we report that LINE1 plays essential roles in mouse embryonic stem cells (ESCs) and pre-implantation embryos. In ESCs, LINE1 acts as a nuclear RNA scaffold that recruits Nucleolin and Kap1/Trim28 to repress Dux, the master activator of a transcriptional program specific to the 2-cell embryo. In parallel, LINE1 RNA mediates binding of Nucleolin and Kap1 to rDNA, promoting rRNA synthesis and ESC self-renewal. In embryos, LINE1 RNA is required for Dux silencing, synthesis of rRNA, and exit from the 2-cell stage. The results reveal an essential partnership between LINE1 RNA, Nucleolin, Kap1, and peri-nucleolar chromatin in the regulation of transcription, developmental potency, and ESC self-renewal.

Keywords:

LINE1, retrotransposons, Nucleolin, Kap1, Dux, MERVL, hypertranscription, rRNA, 2-cell stage, ESCs

Source/Fonte: Cell

+++++

Professores, pesquisadores e alunos de universidades públicas e privadas com acesso ao site Portal de Periódicos CAPES/MEC podem ler gratuitamente este artigo e mais 33.000 publicações científicas.